Showing posts with label Genetic Disorders. Show all posts

Thursday, 3 April 2014

Huntington's Disease by Kivonika Uthayaseelan


                                     Huntington's Disease
                                              By Kivonika Uthayaseelan


                         We learned about the genetics in the previous unit and also how we can have the genetic disorder due to the gene from our parents. Today I will be discussing about my topic and how genetics play an important role with the diseases. So we will be talking about Huntington's disease, which is a neurodegenerative genetic disorder(loss of structure or function of neurons) that affects the muscle coordination. It affects the patient mentally and leads to disorder of losing their action of acquiring knowledge, ability of understanding, and the senses.
                      As we know each of us have total of 46 chromosomes, which come in pairs and one pair from each of the parent. Therefore, we get 23 chromosomes from one parent and the other 23 chromosomes from the other parent. Moreover, there are two types of chromosomes: Autosomal chromosomes and sexual chromosomes.The gene that produces Huntington's disease lies on chromosome 4 and that leads to autosomal dominant, which means the 23rd pair of chromosome (sexual) from a paternal or maternal are not affected. The DNA pattern of Huntington's disease are CAG (Cytosine-Adenine-Guanine) that are repeated over and over again. Individuals with the disorder may have from 40 to over 100 reputation of CAG patterns of DNA.

                     Huntington disease, mostly affects people around 30s to 50s, but in some cases it affects people under the age of 20 and it is called Juvenile Huntington disease. This disorder can affect both genders. This genetic disorder affects people at the age of 30s to 50s due to the uncontrolled muscular movement.

                    The Huntington disease will be diagnosed if there is any changes in the movement of the joints, and also in the emotional and mental state. If anyone notices these changes they have to see the doctor to diagnose the problem. The doctor uses few diagnostic tests to make sure if a patient has Huntington disease. Pre-symptomatic test is the first diagnostic test and this test is allowed to do when a patient doesn't experience any symptoms of Huntington disease.This test will find if a patient has a chance to develop the disorder. The other diagnostic test is called Antenatal test. This test will carry out during pregnancy to make sure the unborn child will not develop Huntington disease. Antenatal test is allowed to the couples, who has a family history of Huntington disease. 

                    The Huntington disease will be detected by the DNA pattern in our body. If a person has 10-20 repeated DNA patterns of CAG, which shows that he or she doesn't have Huntington disease.If a person has 26 CAG patterns of DNA, it will be normal, but more than that it will cause problem of Huntington diseaseAn example of the genetic screening of parents is if a child's grandparent had Huntington disease, then the child 25% more likely the chance to inherit the gene of disorder.


 The symptoms of Huntington disease affect people in three main ways. First in involuntary movement, which means
a person experiences less control in some of the voluntary movement. Sometimes people with the disorder will find it hard to move such as eye movement, loss of balance, difficulty in swallowing,and also difficulty with speaking. Therefore, it affects their daily activities and they have to rely on someone to perform everyday activities. Cognitive problems are the other symptoms of the disease.Cognitive means a person's thinking ability. So people with the disease find it hard to think clearly, response to the question and to concentrate. People also experience a lack of flexibility, difficulty learning new things, lack of awareness of one's behavior, and inability to start a task or conversation. Behavioral symptom is the other symptom people experience with this disease. Behavioral symptoms means a person behavior change with the disease. For example, someone with the disease may get a lot of angry and frustrated. The other symptoms in the behavioral are feeling sadness, loss of interest, changes in appetite, feeling guilty, suicidal thought, tiredness and loss of concentration. The estimated life span of individuals after they have been diagnosed with Huntington disorder is 10-30 years.

                             There is no cure for Huntington disease, but there are drugs to help patients to manage the symptoms. There are medicines which help to reduce involuntary movement and to cure the depression. Mood stabilizers and antipsychotic drugs help the patient to swing the emotional disturbance. Counselling also helps the patient and the family to overcome the disease. Dietitian also guide patients to intake calories and nutrients without losing weight. There is no ethical issue concerning the treatment of this disease because they have all kinds of treatment necessary for the patients health. For example, medication for mood stabilizer, involuntary movement and also counselling and dietitian services also available for the patient.

                        
Stem Cell Research: Huntington  disease


                   In my opinion, this disease is related to genetics, so the researches need to find the other ways to change the individual genes which will benefit a patient with a Huntington disease to lead a happy life. I hope the searchers finds a way to organize music treatments which will help the patients to relax and change their mood.

Question to think about: " Do you think its possible to change the DNA pattern which will beneficial for the patients with Huntington disease? "

References: 
http://www.ncbi.nlm.nih.gov/pubmedhealth/PMH0001775/
http://www.mayoclinic.org/diseases-conditions/huntingtons-disease/basics/definition/con-20030685
http://chealth.canoe.ca/condition_info_details.asp?disease_id=72
https://www.alz.org/dementia/huntingtons-disease-symptoms.asp
http://www.helpguide.org/elder/huntingtons_disease.htm


Phenylketonuria by Kiranjit Singh

Kiranjit Singh                                                                                                                      April 4, 2014
PHENYLKETONURIA
Phenylketonuria, also known as PKU is an autosomal recessive disorder. PKU is a rare disorder. Only one out of fifteen thousand people in the US are born with it. PKU is an inherited disease which means that it is passed down through generations. This disorder increases the levels of phenylalanine in the blood. Phenylalanine is an amino acid, also known as a building block of proteins that is taken in through the diet that the person eats.

Proteins are essential to our body. We need them to repair tissues and make new cells. Amino acids are 20 different building blocks that are made of protein. Protein comes from fish, meat, eggs, nuts and beans that everyone eats. Our bodies are designed to breakdown those proteins into amino acids. Any of the amino acids that are not essential to the body are disposed and are released through the urine. One of the amino acids is called phenylalanine. Phenylalanine cannot process through the bodies of people who are diagnosed with PKU. If phenylalanine cannot be processed, it will cause a buildup in the blood. This buildup will affect your nerve cells and that causes brain damage.


The only way PKU can occur is when the child is born with the disease. The person most likely affected is the child. Plus there is no other way an individual can inherit this disease unless they are born with it. In order for the child to be affected, both parents have to have the recessive-diseased allele. Even if both parents do not have PKU, they can still be carriers of the disease if they are heterozygous. If both parents are heterozygous then there is a 25% chance that their baby might have the disease. Nonetheless, if the child is affected, then that means that the child is homozygous recessive. Furthermore, even if one parent is the carrier of the disease (heterozygous) and the other parent has the disease (homozygous recessive) then that means that there is a 50% chance that the baby will be normal and a 50% chance that the baby will have the disease.

Babies are born with PKU. As soon as the baby is born, it is the doctor’s job to test the baby for PKU with a simple blood test. The reason only newborn babies are affected by this disease is depending on if the parents have the diseased allele. If the parents do not have the diseased allele then there is no possibility that the child will be diagnosed with that disease.

If PKU is not treated early, as in if the parents are unaware that their baby had this disease, what will happen is that the first few months the baby will seem to be normal but after that time period you will notice the symptoms. You will notice that the baby has started to lose interest in things when they are 3 to 7 months old. When the baby is one year old, it starts to become obvious that the baby’s brain is not developing the way a normal baby’s brain would develop. Other symptoms include dry skin, seizures, rash or the baby starting to get smelly because the phenylalanine is in their urine, breath and sweat. Often, kids with untreated PKU have small heads called microcephaly. This means that if this disease is not treated on time then the phenylalanine can build up into harmful levels in the body. This can cause permanent mental retardation along with other health problems.

Thankfully, PKU is a treatable disease. The treatment involves the person to have a diet with low in phenylalanine, preferably when the child is growing. It is important that this diet be strictly followed. A person with PKU can live a normal healthy life span with a normal mentality if they continue on with this diet for the rest of their lives. The only problem there can be in the treatment of this disease is if this disease is not caught on time. If this is the case then the individual can suffer through permanent intellectual disability.

Do you think people who are diagnosed with PKU should have children and risk causing their disease to pass onto their children? Why or why not?

References:


Wednesday, 2 April 2014

Neurofibromatosis By: Gurneet Bhela


Neurofibromatosis (NF)
____________________________________


BY: Gurneet Bhela
Neurofibromatosis is autosomal dominant genetic disorder that affects the nervous system also including the brain, spinal cord, nerves, skin, and other body systems in the body. People affected by this disorder develop non-cancerous tumours along their nerves, nerve tissue and under the skin. The non-cancerous tumours formed are called neurofibromas which disturb cell growth within the nervous system. There are two types of Neurofibromatosis- NF1 and NF2. The severity of both types of neurofibromatosis varies greatly. NF1 is more common and the chromosome that is affected is chromosome 17. NF-1 is caused by a mutation of a gene on the long arm of chromosome 17 which encodes a protein known as neurofibromin which plays an important role in cell signaling. For NF2 chromosome 22 is affected. NF2 is caused by mutations of the "Merlin" gene which influences the form and movement of cells.


Neurofibromatosis is an inherited disorder therefore an affected person has 1 chance in 2 of passing it on with each pregnancy but in some cases Neurofibromatosis can actually be the result of a spontaneous change (mutation) in the genetic material of the sperm or egg at conception even in families with no previous history of Neurofibromatosis.

A doctor who treats NF1, such as a neurologist, geneticist, dermatologist, or developmental pediatrician will diagnose this condition. NF1 is can sometimes be diagnosed in younger children who appear to have certain symptoms of spots on the skin that are at least ½ inch in size. But the spots are not enough for a complete diagnosis so when the grow continues to grow at the age of 10 Neurofibromas will begin to become evident on various parts of the body, beginning with the arms and then spreading to other parts.
For a child to be diagnosed with NF1 he/she must have at least two of the following symptons:
  • café-au-lait spots of a certain number, size, and location
  •  the appearance of two or more neurofibromas
  •  an optic glioma (tumor along the main nerve of the eye that is responsible for sight)
  •  certain skeletal abnormalities
  •  a family member with NF1
  • freckling under the arms or in the groin


café-au-lait spot

NF2 is usually not diagnosed until a child is older. Hearing loss in the late teens and early twenties is often among the first symptoms of the disorder, and is caused by tumors growing on the auditory nerves (which carry electrical impulses from the inner ear to the brain).
Tests like magnetic resonance imaging (MRI) and X-rays may be used to screen for tumors or evidence of skeletal problems. A child's head circumference will be measured because kids with symptoms of NF can have a circumference that's larger than normal for their age.

To diagnose NF2, doctors will check for any evidence of hearing loss. Audiometry (hearing tests) can be used as well as imaging tests to look for tumors in the nerves of the ears, spinal cord, or brain.

Genetic testing is available for people with a family history of either NF1 or NF2. Although testing is still not 100% sensitive, recent advances have increased sensitivity to over 90%. Prenatal testing may be used to identify the disorder of NF-1 in the fetus. Preimplantation genetic diagnosis and Chorionic villus sampling can be used to screen for Neurofibromatosis.

Preimplantation genetic diagnosis
NOTE: Preimplantation genetic diagnosis: a technique using in vitro fertilization to ensure that a baby does not possess a known genetic defect of either parent
Chorionic villus sampling: is a form of prenatal diagnosis to determine chromosomal or genetic disorders in the fetus


People diagnosed with Neurofibromatosis have a normal life span.  Neurofibromas on or under the skin can increase with age and cause cosmetic and psychological issues.
Once a person has developed Neurofibromatosis there is no cure that can completely reverse the genetic disorder. Instead the treatments for neurofibromatosis focus on controlling the existing symptoms. For a more complex case where the symptoms are high and dangerous, options for treatment are necessary. Surgery can be performed to remove problematic growths or tumors, chemotherapy or radiation if a tumor has turned malignant or cancerous, surgery for bone problems and therapy. These treatments can be used for both NF1 and NF2 depending on the individual’s progress. Although this treatment is not able to cure the disease, it is still possible to live a normal healthy life if the symptoms are regularly monitored and controlled so it is important to be diagnosed at an earlier age.

There are no ethical issues concerning the treatment of this disease but it is a rare genetic that deserves to raise awareness for. Recently, non-profit and charitable organizations have started to bring awareness and education about Neurofibromatoses to mainstream media in order to better connect those affected with helpful resources to lead more productive and satisfying lives.


In terms of further studies regarding Neurofibromatosis, there are many organizations who have the goal to help provide better therapies for affected patients, provide outstanding clinical care and also educating families about this genetic disorder. I believe Neurofibromatosis is a more complicated disease because it is concerned with the nervous system which makes it more difficult to study the body’s nerves. But the focus of study should be directed towards how this mutation occurs in the chromosomes and how it changes the genes of a person’s appearance very rapidly. There is a need to study the cause rather than the result; further studies need to be targeted to understand which changes in the NF1 gene that leads to various symptoms and associated features. The two chromosomes are connected to the function of the brain and nervous system so perhaps studying how brain productivity and function changes and to what scale compared to a healthy human body will be useful for finding a potential cure.


                                                           A question for thought:

Why do you think the symptoms for NF1 and NF2 are quite different from each other even though they are both a genetic disorder of Neurofibromatosis? If you were a scientist, doctor or researcher which type of Neurofibromatosis do you think would be more important to study further NF1 or NF2?


References:


Tuesday, 1 April 2014

Hemophilia: By: Zainab Manjra


                                                Hemophilia

      We all sort of have a basic idea on what hemophilia is but do we really know the whole background of it and how it occurs? Read on to find out!

  Hemophilia is basically a group of hereditary genetic disorder that damages the body's ability to control thee blood clotting, which is used to stop bleeding when the blood vessel is broken. The most common form of the disorder, that is present in about 1 in 5,000-10,000 male births, is in fact Hemophilia. It occurs in around 1 in about 20,000-34,000 male births. Hemophilia affects the X chromosomes and the disorder generally affects the males because, women are the carriers of the disease and since they have an additional X chromosome to cancel out the bad gene they are not affected. Men who have hemophilia, they cannot pass down the disorder to their sons because they would be passing the Y chromosome, which is healthy, but the daughters would be carriers of the disease since they are receiving the fathers affected X chromosome. Thus hemophilia is considered a sex-linked disease.

   What causes hemophilia? Hemophilia is caused but genetic makeup but it is also possible for someone to acquire hemophilia through a unplanned genetic mutation. It can also be developed if the body forms antibodies to clotting factors in the blood that prohibit the clotting factors from working.

   Hemophilia can be detected by going to your family doctor who will ask questions about your personal and family medical history. If this patient does not have any family or personal medical history then physical exam and blood tests may be done to diagnose hemophilia. the blood tests are used to find out the following:  
  • How long it takes for your blood to clot
  • Whether your blood has low levels of any clotting factors
  • Whether any clotting factors are completely missing from  your blood
The blood tests will identify whether or not you have hemophilia, how severe it is and what type of hemophilia it is. The estimated lifespan of a hemophiliac is about 50-60 years.

The following are some symptoms of hemophilia:
  • Bleeding into joints 
  • Bleeding into soft tissues and muscles
  • Bleeding in the mouth from a cut, bitten tongue etc
  • Blood in the urine
  • Surface bruising
There are many ways to treat hemophilia:
  • Replacement therapy ( In which they infect concentrates of clotting factor VIII, helps replace clotting factor that is running low or missing)
  • Desmopressin
  • Antifibrinolytic Medicines 
  • Gene therapy
  • Treatment of a specific bleeding site
Question: Do you think Hemophiliacs should play rough sports outside? Why or why not?


List of References:

Saturday, 29 March 2014

Alkaptonuria By: Saidia Ali

Alkaptonuria

By: Saidia Ali 

We are currently studying the genetics unit and we all know the basics now. In the beginning of this unit we learned about karyotypes and non-disjunction. All the information related to that will be useful in understanding this disorder better. So, get ready to learn about alkaptonuria! 

Alkaptonuria also called AKU and black urine disease  is a very rare genetic disorder that occurs because of a mutation in chromosome 3. Alkaptonuria is an inherited recessive disorder which means that it must be passed on by both parents. People who have one copy of the gene are carriers of the disease and do not suffer from it but there is a twenty-five percent chance of them passing it to their offspring. We know this because we have done Punnett square problems related to similar situations. 

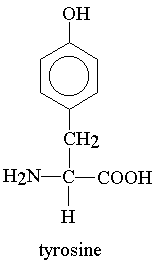
Now that we know some basic information about the inheritance of alkaptonuria but what exactly is it? Alkaptonuria is a disorder caused by the lack of an enzyme homogentisic dioxygenase (HGD). Patients who have alkaptonuria are unable to break down a substance called homogentistic acid. As a result this acid starts to build up in the body leading to many severe problems which we will discuss later on. Patients with this disease are also unable to metabolize the amino acids called phenylalanine and tyrosine. These two amino acids are important in producing hormones and pigments. As I've mentioned before, alkaptonuria is a rare disease and it is estimated that it affects one out of 200,000 worldwide. In some countries AKU is more common like in Slovakia and the Dominican Republic where it affects one in every 19,000. AKU is not a disease that is only inherited by men or women, it is a disease that affects men, women, and children equally. AKU patients have a normal life expectancy. 


Sometimes there are diseases where the patients cannot feel or see the symptoms but a patient with AKU can see the problems. Here is a list of the symptoms: 

  • One of the earliest signs in a baby are dark stains in his/her diaper.
  • Urine turning black or brown when exposed to air is a common symptom
  • Stained cartilage in the ear 
  • Darkening of skin
  • Arthritis in knees, back, and/or hips
The buildup of HGD can lead to heart problems. 


When AKU is suspected here are ways to diagnose the patient. One way is gas chromatography which is used to check for homogentisic acid in the urine. DNA testing can also be done to look for the mutated gene.
 Although there is no particular treatment for AKU there are ways to lower the symptoms and keep the disease from getting out of control. Many doctors recommend exercising and physiotherapy to assist the patient with arthritis. Maintaining a healthy diet is also key. Evidence shows that increasing the input of vitamin C can decrease the development of arthritis. Another crucial change in an AKU patient's diet is eliminating food that contain phenylalanine and tyrosine. These foods include meats, soy, nuts and seeds, and artificial sweeteners. 

Learning about this topic was very interesting and I hope that you enjoyed learning about this topic just as much as I did researching it.

Question: Do you think genetic engineering will help us find a cure for alkaptonuria in the future? Why or why not?


http://www.ncbi.nlm.nih.gov/pmc/articles/PMC3108520/